FDA Medical Device Testing: Key Updates for 2026
Anúncios
Understanding the significant changes in FDA medical device testing regulations for 2026 is crucial for manufacturers to ensure compliance, accelerate market entry, and uphold the highest standards of patient safety and product efficacy.
The medical device industry is a dynamic and highly regulated sector, constantly evolving to meet new technological advancements and address emerging public health needs. For manufacturers, staying abreast of regulatory changes is not just about compliance; it’s about safeguarding patient well-being and ensuring market viability. The year 2026 brings forth significant updates in how the U.S. Food and Drug Administration (FDA) approaches medical device testing, demanding proactive engagement and strategic adaptation from all stakeholders. This article delves into the critical shifts, offering a comprehensive guide to navigating the revised landscape.
Anúncios
Understanding the Evolving FDA Landscape for 2026
The FDA’s regulatory framework for medical devices is designed to ensure their safety and effectiveness. As technology advances and medical science progresses, these regulations must evolve to keep pace. The updates slated for 2026 reflect the agency’s commitment to fostering innovation while maintaining stringent oversight. Manufacturers must recognize that these aren’t minor tweaks but rather foundational adjustments that will influence every stage of device development and approval.
Anúncios
These changes are often driven by several factors, including lessons learned from past device performance, the emergence of new technologies like AI and machine learning in medical devices, and global harmonization efforts. Understanding the underlying philosophy behind these shifts can help companies better prepare and integrate new requirements into their existing quality management systems.
Key Drivers Behind Regulatory Shifts
- Technological Advancements: The rapid integration of AI, machine learning, and digital health technologies into medical devices necessitates new testing paradigms.
- Patient Safety Enhancements: Continuous review of adverse event reports and post-market surveillance data informs regulatory adjustments aimed at improving patient outcomes.
- Global Harmonization: Efforts to align FDA regulations with international standards, such as those from the International Medical Device Regulators Forum (IMDRF), streamline global market access.
- Supply Chain Resilience: Recent global events have highlighted the need for more robust testing and oversight of supply chains to ensure device quality and availability.
In essence, the 2026 updates represent a concerted effort by the FDA to modernize its regulatory approach, making it more responsive to innovation and more effective in protecting public health. Companies that embrace these changes early will gain a significant competitive advantage.
New Pre-Market Submission Requirements and Testing Protocols
One of the most impactful areas of change for 2026 lies within pre-market submission requirements and the associated testing protocols. The FDA is increasingly emphasizing a data-driven approach, requiring more comprehensive and robust evidence of device performance and safety before market authorization. This includes a heightened focus on real-world evidence and a more detailed demonstration of device functionality across diverse patient populations.
Manufacturers should anticipate a need for more sophisticated testing methodologies, potentially involving advanced simulations, in silico trials, and expanded clinical investigations. The goal is to minimize uncertainty about a device’s performance in actual use conditions. This shift necessitates a re-evaluation of current testing strategies and an investment in new capabilities.
Enhanced Clinical Data Expectations
The FDA is moving towards requiring more granular and representative clinical data. This means studies may need to include a broader range of demographics, comorbidities, and use cases to reflect the real-world patient population more accurately. The agency is also keen on understanding long-term efficacy and safety profiles, which might influence the design of post-market studies.
- Diversity in Trial Participants: Greater emphasis on including diverse ethnic, age, and gender groups in clinical trials.
- Real-World Data Integration: Encouragement of real-world evidence (RWE) to supplement traditional clinical trial data, particularly for post-market surveillance and label expansion.
- Patient-Reported Outcomes (PROs): Increased focus on collecting and analyzing PROs to understand the patient’s perspective on device benefit and risk.
These enhanced expectations for clinical data underscore the importance of meticulous study design and execution, ensuring that the collected evidence directly addresses the FDA’s safety and effectiveness concerns.
Cybersecurity in Medical Devices: A Critical Testing Frontier
As medical devices become increasingly interconnected and reliant on software, cybersecurity has emerged as a paramount concern for the FDA. The 2026 updates place a significantly stronger emphasis on the cybersecurity robustness of devices, recognizing that vulnerabilities can pose direct threats to patient safety and data privacy. Manufacturers are now expected to integrate cybersecurity considerations throughout the entire device lifecycle, from design to post-market monitoring.
Testing protocols will need to include rigorous assessments of a device’s resilience against cyber threats, its ability to maintain essential functions during an attack, and its mechanisms for patching vulnerabilities. This is not merely an IT concern; it is a fundamental aspect of device safety and efficacy that requires specialized expertise.
Mandatory Cybersecurity Testing Components
Manufacturers should prepare for mandatory cybersecurity testing components within their submission packages. These will likely include:
- Threat Modeling: Comprehensive analysis to identify potential cybersecurity risks and vulnerabilities.
- Vulnerability Assessments: Testing to identify and quantify security weaknesses in the device’s software and hardware.
- Penetration Testing: Simulating cyberattacks to evaluate the device’s defenses and response capabilities.
- Software Bill of Materials (SBOM): Providing a detailed list of all software components, including open-source elements, to facilitate vulnerability tracking.
Ensuring robust cybersecurity is no longer an optional add-on but a core requirement for medical device approval. Companies must invest in specialized cybersecurity expertise and integrate these considerations into their product development from the earliest stages.
The Role of Digital Health Technologies and AI in Testing
The proliferation of digital health technologies (DHTs) and artificial intelligence (AI) in medical devices presents both immense opportunities and unique regulatory challenges. The FDA’s 2026 updates aim to provide clearer pathways for the review and testing of these innovative technologies. This involves developing new guidance documents and potentially new regulatory classifications that specifically address the unique characteristics of AI/ML-driven devices, such as their adaptive algorithms and continuous learning capabilities.
Testing for AI-enabled devices goes beyond traditional hardware and software validation. It requires demonstrating the AI’s performance across various datasets, assessing its fairness and bias, and establishing clear mechanisms for monitoring its behavior post-market. Manufacturers must be prepared to provide extensive documentation on their AI models, training data, and validation processes.
Specific Considerations for AI/ML Devices
- Algorithm Transparency: Requirements for detailed documentation of AI algorithms, including how decisions are made and potential sources of bias.
- Data Governance: Emphasis on the quality, representativeness, and security of the data used to train and validate AI models.
- Continuous Learning Plans: For adaptive AI, clear plans for how the device will be monitored, updated, and re-validated post-market.
- Performance Metrics: Establishing appropriate and measurable performance metrics for AI algorithms, especially in dynamic clinical environments.
The FDA is keen to support innovation in DHTs and AI but will do so with a strong focus on ensuring these technologies are safe, effective, and ethically deployed. Proactive engagement with FDA guidance and pilot programs can be highly beneficial for manufacturers in this space.
Post-Market Surveillance and Real-World Evidence
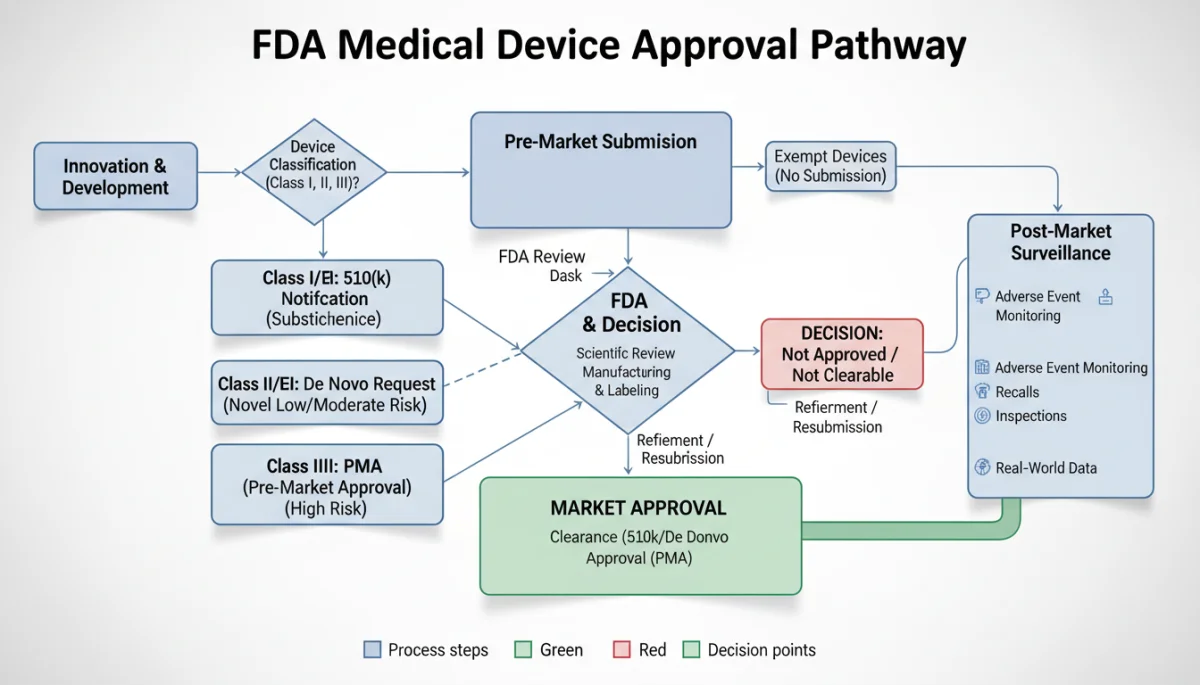
While pre-market testing ensures a device’s safety and efficacy before it reaches patients, post-market surveillance (PMS) is equally critical for monitoring its performance in real-world settings. The 2026 FDA updates are expected to strengthen PMS requirements, leveraging real-world evidence (RWE) more extensively to detect unforeseen issues, evaluate long-term outcomes, and inform regulatory actions. This shift emphasizes a continuous lifecycle approach to device regulation.
Manufacturers will need to develop more robust PMS plans, potentially integrating data from electronic health records, patient registries, and wearable devices. The ability to collect, analyze, and report RWE effectively will become a cornerstone of ongoing compliance. This proactive monitoring helps identify trends, refine device usage instructions, and even prompt recalls or design modifications when necessary.
Key Aspects of Enhanced Post-Market Surveillance
The FDA’s updated approach to PMS will likely include:
- Active Surveillance Systems: Utilizing advanced analytics and potentially AI-driven tools to actively monitor device performance and identify adverse events.
- Real-World Data Integration: Greater reliance on RWD for ongoing risk-benefit assessments and to support labeling changes or expanded indications.
- Manufacturer Reporting Obligations: Clearer and potentially more frequent reporting requirements for adverse events and device malfunctions.
- Patient Engagement: Encouragement of mechanisms for direct patient feedback and reporting of device experiences.
A comprehensive and well-executed PMS strategy is vital not only for regulatory compliance but also for building trust with both the FDA and the end-users of medical devices. It allows for continuous improvement and rapid response to any emerging safety signals.
Strategies for Successful Compliance in 2026
Navigating the complex and evolving landscape of FDA regulations for medical device testing in 2026 requires a strategic and proactive approach. Manufacturers cannot afford to wait until the last minute to adapt; early preparation and a thorough understanding of the new requirements are paramount. This involves not only updating internal processes but also fostering a culture of continuous learning and regulatory intelligence within the organization.
Developing a robust regulatory strategy that anticipates future changes, invests in necessary testing capabilities, and engages with the FDA early can significantly streamline the approval process and ensure sustained market access. Compliance should be viewed not as a burden, but as an integral part of product quality and patient safety.
Actionable Steps for Manufacturers
- Conduct a Gap Analysis: Assess current testing protocols and quality management systems against the anticipated 2026 FDA requirements to identify areas needing improvement.
- Invest in New Technologies: Adopt advanced testing methodologies, cybersecurity tools, and data analytics platforms to meet enhanced data expectations.
- Enhance Internal Expertise: Train regulatory affairs, R&D, and quality assurance teams on the new guidelines, potentially hiring specialists in areas like cybersecurity or AI validation.
- Engage with the FDA: Utilize pre-submission meetings and participate in pilot programs or workshops to gain clarity and align strategies with the agency’s expectations.
- Strengthen Supply Chain Oversight: Implement more rigorous testing and audit procedures for suppliers to ensure component quality and reliability.
By taking these steps, medical device manufacturers can transform regulatory challenges into opportunities for innovation and ensure their products continue to meet the highest standards of safety and effectiveness.
| Key Regulatory Area | Brief Description of 2026 Updates |
|---|---|
| Pre-Market Submissions | Increased emphasis on robust clinical data, real-world evidence, and comprehensive performance testing for new devices. |
| Cybersecurity | Mandatory integration of cybersecurity testing, threat modeling, and vulnerability assessments throughout the device lifecycle. |
| Digital Health & AI | New guidance for AI/ML device testing, focusing on algorithm transparency, data governance, and continuous learning plans. |
| Post-Market Surveillance | Enhanced requirements for active surveillance, real-world evidence integration, and more frequent reporting. |
Frequently Asked Questions About FDA Medical Device Testing in 2026
The biggest changes involve enhanced pre-market data requirements, mandatory cybersecurity testing, specific guidance for AI/ML devices, and strengthened post-market surveillance using real-world evidence. These updates aim to modernize oversight and adapt to technological advancements.
Manufacturers must now integrate robust cybersecurity testing, including threat modeling, vulnerability assessments, and penetration testing, into their development cycles. This ensures devices are resilient against cyber threats and patient data remains secure, affecting design and validation processes.
The FDA is developing clearer pathways for AI/ML devices, focusing on algorithm transparency, data governance, and continuous learning plans. Testing will need to demonstrate the AI’s performance across diverse datasets and establish clear post-market monitoring mechanisms.
Real-world evidence provides crucial insights into a device’s performance in actual clinical use, complementing traditional trial data. The FDA uses RWE to assess long-term safety and efficacy, inform labeling changes, and enhance post-market surveillance, ensuring continuous patient safety.
Manufacturers should conduct a gap analysis, invest in new testing technologies, enhance internal expertise in areas like cybersecurity and AI, and engage proactively with the FDA through pre-submission meetings. Early preparation is key to successful compliance and market access.
Conclusion
The upcoming FDA regulatory updates for medical device testing in 2026 mark a significant evolution in how medical devices are evaluated, approved, and monitored. These changes reflect a commitment to patient safety, technological adaptation, and global harmonization. For manufacturers, understanding and proactively addressing these shifts is not merely a compliance exercise but a strategic imperative. By investing in advanced testing methodologies, strengthening cybersecurity, embracing digital health guidance, and leveraging real-world evidence, companies can navigate this evolving landscape successfully, ensuring their innovative devices continue to reach patients safely and efficiently.
